UK Market Compliance Support
Market planning that distinguishes Great Britain from Northern Ireland and accounts for current CE transition rules.
Quick Consultation
Market planning that distinguishes Great Britain from Northern Ireland and accounts for current CE transition rules.
Quick Consultation
Registration or listing does not mean FDA approval, clearance or authorization.
Support for applicable establishment registration and device listing under 21 CFR Part 807.
Industrial safety, medical distribution, and e-commerce channels.
CE marking support under the applicable medical-device or PPE framework, determined by each component.
Quick Consultation
Identify whether components fall under Regulation (EU) 2017/745, the PPE Regulation or another applicable framework.
Manufacturing under an ISO 13485-certified medical-device quality management system.
Multilingual label and instruction coordination, including manufacturer, importer or authorized-representative details where required.
Market planning that distinguishes Great Britain from Northern Ireland and accounts for current CE transition rules.
Quick Consultation
Support for applicable MHRA registration, UK Responsible Person information and valid marking route. CE-marked devices remain recognized in Great Britain during current transition periods, depending on device and legislation.
CE or CE UKNI requirements apply under the separate Northern Ireland framework; UKCA alone is not recognized.
Workplace first aid kit configuration support as a British standard reference, not a substitute for medical-device market authorization.
A risk-class pathway separating establishment licensing from product licensing.
Quick Consultation
Medical Device Establishment Licence coordination for Class I manufacturers and importers or distributors of devices across classes. An MDEL does not approve individual products.
Medical Device Licence file support for Class II, III or IV devices where required.
English and French label and information coordination according to applicable requirements.
SFDA market-access support coordinated with the responsible local economic operator.
Quick Consultation
Configuration-specific review for the applicable SFDA medical-device marketing authorization route.
Document support for local representative, importer and establishment requirements where applicable.
Arabic information, labels, certificates and shipment-document coordination based on the approved route.
ARTG inclusion support through an Australian-based sponsor for applicable medical devices.
Quick Consultation
The sponsor must be an Australian-based legal entity and holds defined regulatory responsibilities.
Prepare manufacturer and device information for inclusion in the Australian Register of Therapeutic Goods where required.
Support evidence, declarations, labels and instructions against classification-specific requirements.
NMPA filing or registration support based on device classification and product-specific requirements.
Quick Consultation
Determine whether individual products are Class I, II or III medical devices under the current catalog and rules.
Class I filing or Class II–III registration documentation support where applicable.
Chinese labeling, instructions, technical files and domestic responsible-party coordination.
A controlled documentation core adapted into market-specific submission, label and packaging sets.
Quick Consultation
Compare target-market requirements against the exact product and component file set.
Organize available declarations, reports, specifications, labels and production records by SKU.
Maintain separate approved versions for language, responsible parties, claims, identifiers and shipping marks.
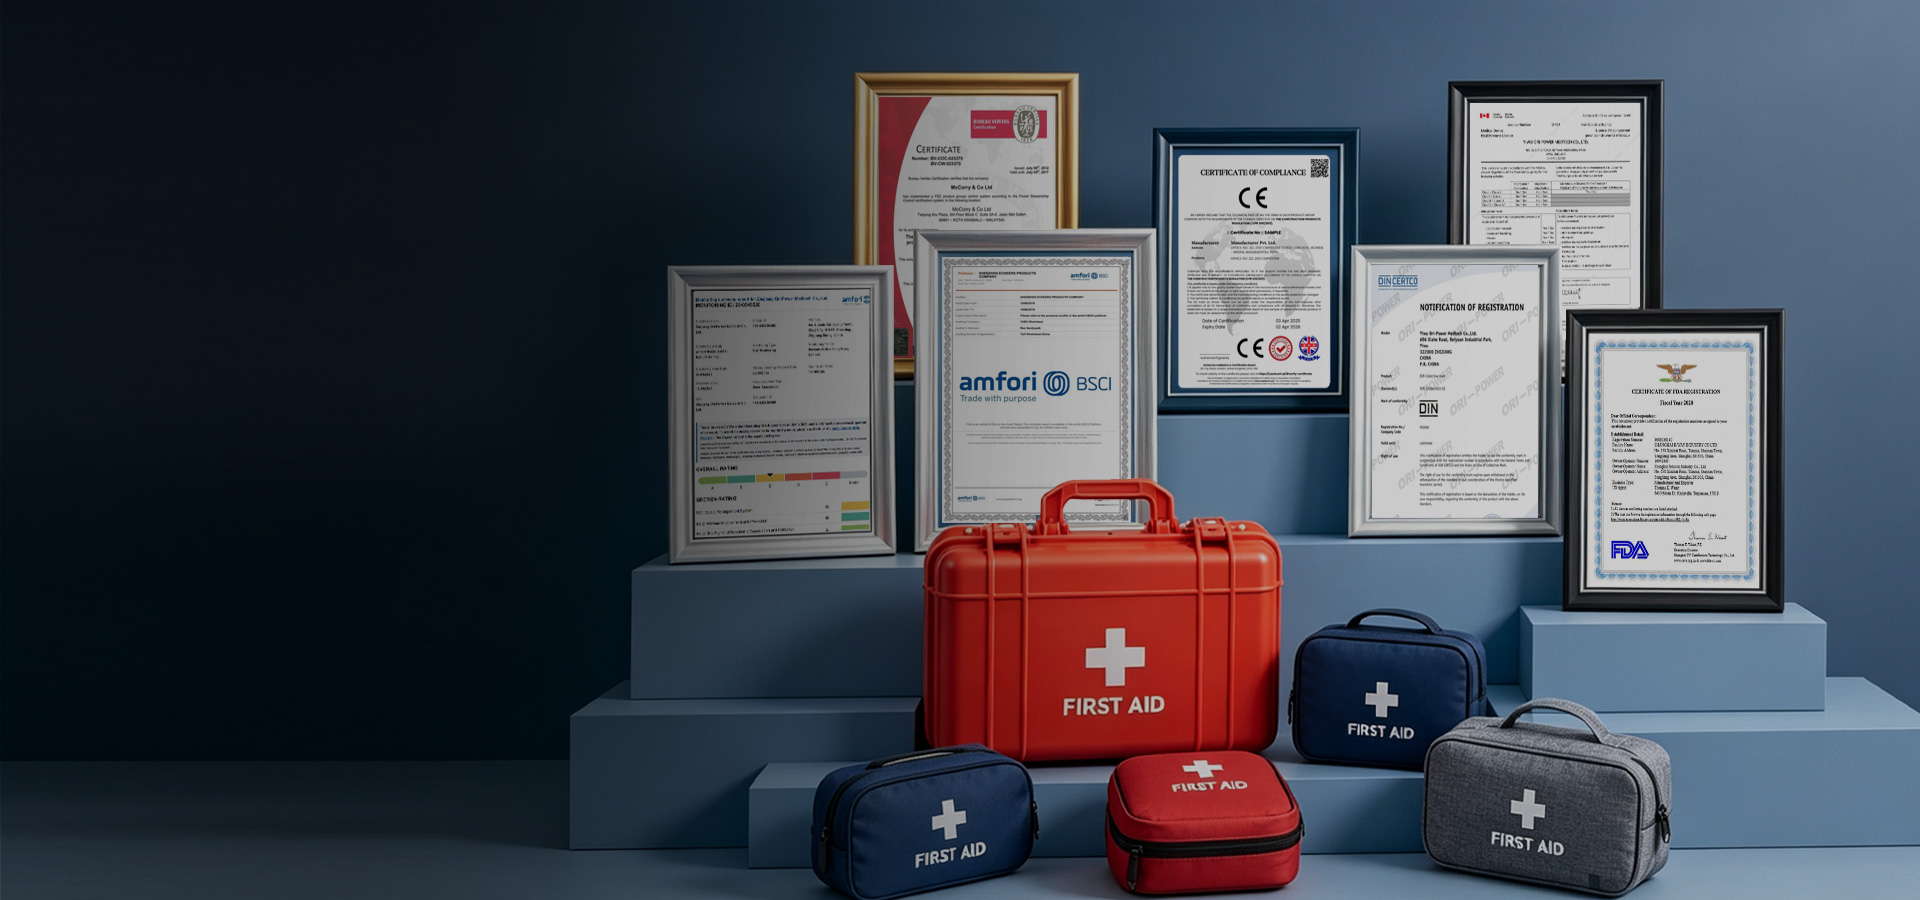
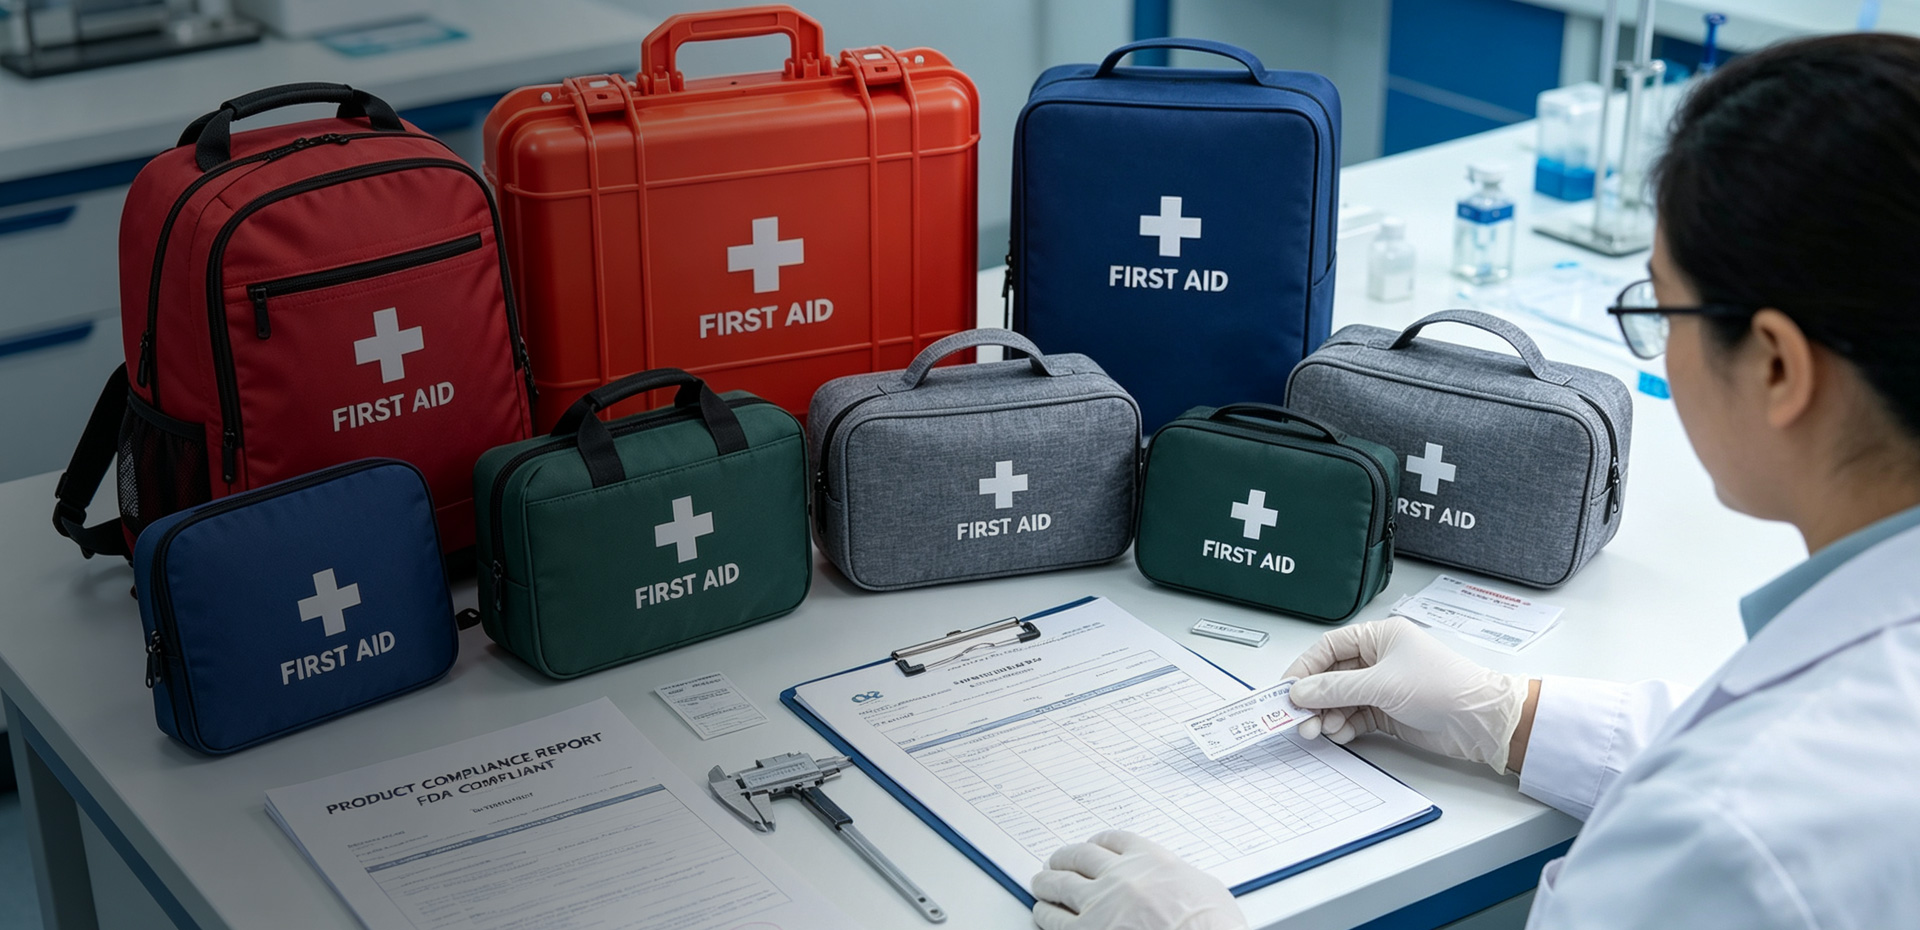
CE marking, FDA, and market-specific documentation support
Plan required operator, importer, warning, language, and product identification information.
Coordinate outer packaging, instructions for use and retail presentation without breaking label hierarchy.
Identify missing evidence, market-representative needs and import-document dependencies early.
Use established workplace or vehicle-kit references where commercially appropriate, while keeping standards distinct from regulatory authorization.
Device-identifier data preparation and label coordination where the applicable market and device require it.
Lot-level identification to support receiving, distribution, complaint handling, and recall readiness.
Controlled manufacturing and inspection records linked to finished product batches.
Version-controlled artwork, language, barcode, and outer carton information.
Configuration-specific documentation packages assembled from available declarations, reports, and product evidence.
Support for importer, representative, sponsor or registration submissions according to the target route.

+86 18507974886